
11.3 Structures of Solids



PRE-HEALTH PREP


Structures of Solids
Cubic Unit Cells and Close Packing
General Chemistry 11.3
Lesson Overview
Crystalline solids organize into repeating unit cells, with the three cubic types—simple cubic, body-centered cubic, and face-centered cubic—differing in atom count, coordination number, and packing efficiency (52%, 68%, and 74%, respectively). Close-packed structures achieve optimal space filling through either hexagonal (ABAB) or cubic (ABCABC) layer stacking. The four categories of solids—molecular, network covalent, ionic, and metallic—exhibit melting points that reflect the strength of their bonding interactions, from weak intermolecular forces to extended covalent networks.
3 Types of Cubic Unit Cells
Most solids adopt a repeating structural pattern known as a crystal lattice. When the basic repeating unit is a cube, we call it a cubic unit cell. There are three types of cubic unit cells: simple cubic (SC), body-centered cubic (BCC), and face-centered cubic (FCC). Understanding each crystal structure is essential for general chemistry and explains differences in packing efficiency, density, and physical properties of crystalline solids.
What About Amorphous Solids?
Not every solid follows a repeating crystal pattern. Amorphous solids such as glass lack any regular repeating structure. However, the vast majority of solids discussed in this chapter are crystalline and adopt some type of repeating unit cell.
All three cubic unit cell types share one feature: atoms occupy each of the eight corners of the cube. What distinguishes them is where additional atoms appear.
| Unit Cell Type | Atoms at Corners | Additional Atom Positions |
|---|---|---|
| Simple Cubic (SC) | 8 corners | None |
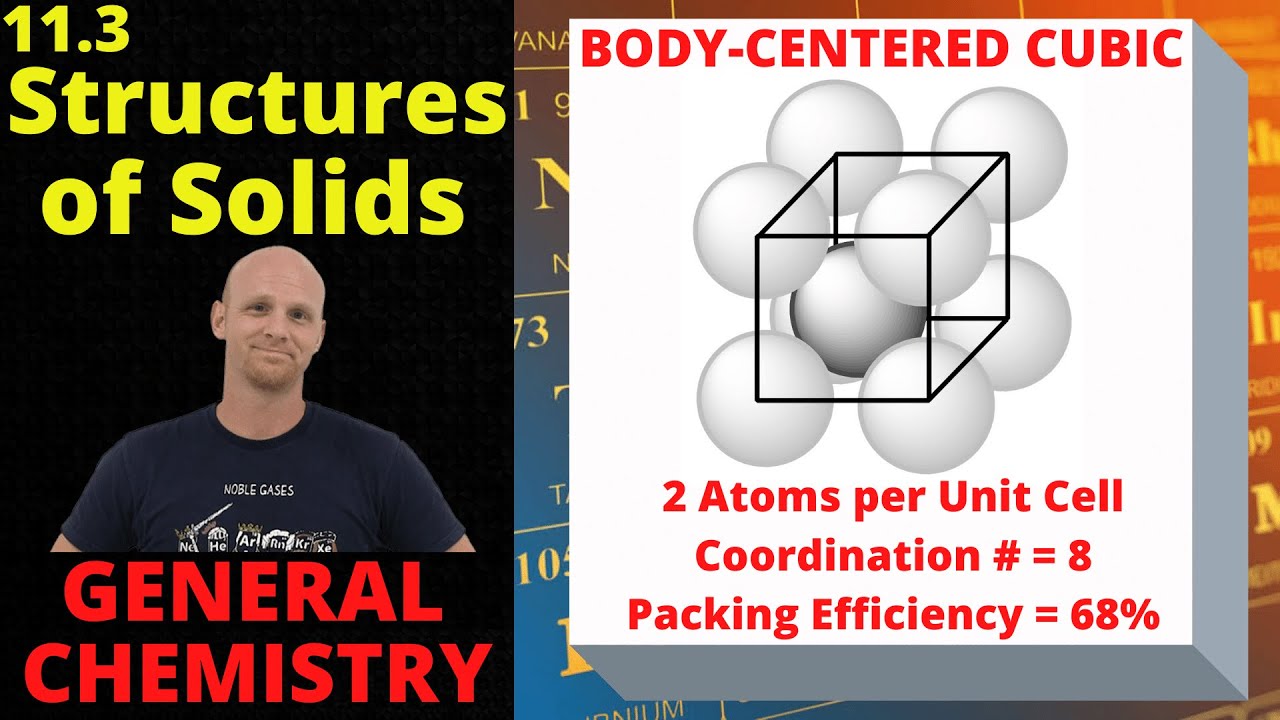
| Body-Centered Cubic (BCC) | 8 corners | 1 atom at the center of the cube |
| Face-Centered Cubic (FCC) | 8 corners | 1 atom at each of the 6 face centers |
In the body-centered cubic structure, the additional atom sits right in the middle of the cube, not touching any edge. In the face-centered cubic structure, six additional atoms appear at the center of each face—top, bottom, left, right, front, and back.

Simple Cubic (SC)

Body-Centered Cubic (BCC)

Face-Centered Cubic (FCC)
How Many Atoms Per Unit Cell?
Atoms at different positions within a unit cell are shared with neighboring unit cells, so each position contributes only a fraction of an atom. Calculating the number of atoms per unit cell is a foundational step in all cubic unit cell problems.
Corner Atoms: Shared 8 Ways
Every corner atom is shared by eight adjacent unit cells—four in the layer below and four in the layer above. Each corner therefore contributes only 1/8 of an atom to any single unit cell.
Face-Center Atoms: Shared 2 Ways
An atom at the center of a face is shared between exactly two unit cells—the one above and the one below that face. Each face-center atom contributes 1/2 of an atom to a single unit cell.
Body-Center Atoms: Not Shared
The atom in the center of a body-centered cubic cell belongs entirely to that unit cell. It contributes 1 full atom.
| Unit Cell Type | Corner Contribution | Additional Contribution | Total Atoms |
|---|---|---|---|
| Simple Cubic | 8 × 1/8 = 1 | None | 1 |
| Body-Centered Cubic | 8 × 1/8 = 1 | 1 body center × 1 = 1 | 2 |
| Face-Centered Cubic | 8 × 1/8 = 1 | 6 face centers × 1/2 = 3 | 4 |
Memorize These
SC = 1 atom, BCC = 2 atoms, FCC = 4 atoms per unit cell. These values appear repeatedly in calculations throughout this chapter.
What Is the Coordination Number in Crystal Structures?
The coordination number is the number of nearest-neighbor atoms surrounding any given atom in the crystal structure. It reflects how tightly packed the atoms are and is a key property that distinguishes the three cubic unit cells.
Simple Cubic: Coordination Number = 6
In the simple cubic structure, each corner atom has six nearest neighbors—one in each direction along the three axes (left, right, front, back, above, below).
Body-Centered Cubic: Coordination Number = 8
The body-center atom is equidistant from all eight corner atoms, giving it eight nearest neighbors. Every atom in a BCC lattice shares this coordination number of 8.
Face-Centered Cubic: Coordination Number = 12
Consider a face-center atom on the top face of the cube. It has four nearest neighbors at the corners of that face, four face-center atoms on the adjacent faces in the layer below, and four more face-center atoms in the layer above. That yields 12 nearest neighbors—the highest coordination number among the cubic unit cells.
| Unit Cell Type | Atoms Per Unit Cell | Coordination Number |
|---|---|---|
| Simple Cubic | 1 | 6 |
| Body-Centered Cubic | 2 | 8 |
| Face-Centered Cubic | 4 | 12 |
How Do Edge Length and Atomic Radius Relate?
In each cubic unit cell, the edge length a relates to the atomic radius r differently, depending on where the atoms actually touch. Knowing these relationships is essential for density calculations and atomic radius problems.
Simple Cubic: Atoms Touch Along the Edge
In the simple cubic structure, adjacent corner atoms touch directly along each edge. The edge spans one radius from the left atom and one radius from the right atom:
Face-Centered Cubic: Atoms Touch Along the Face Diagonal
In the FCC structure, atoms do not touch along the edge. Instead, they touch along the diagonal of each face. Traveling across that diagonal, you pass through one radius (corner atom), one full diameter (face-center atom), and one more radius (far corner atom)—a total of four radii. Using the Pythagorean theorem on a face with sides of length a:
Body-Centered Cubic: Atoms Touch Along the Body Diagonal
The BCC structure is the trickiest. Atoms touch neither along an edge nor along a face diagonal—they touch along the body diagonal, running from one corner through the center atom to the opposite corner. Applying the Pythagorean theorem in three dimensions:
a² + 2a² = c²
c = √3 × a
Along that body diagonal: one radius + one diameter + one radius = 4r, giving:
Exam Tip
Exam problems frequently give you an edge length and ask for the atomic radius (or vice versa). The key is selecting the correct relationship for the unit cell type:
- SC: a = 2r
- FCC: √2 · a = 4r
- BCC: √3 · a = 4r
Packing Efficiency of the 3 Cubic Unit Cells
Packing efficiency measures the percentage of a unit cell's volume that is actually occupied by atoms, as opposed to empty space. Think of a ball pit—no matter how you arrange spheres, gaps remain between them. The same is true at the atomic level in all crystal structures.
The calculation divides the total volume of atoms in the unit cell by the total volume of the cube, then multiplies by 100:
The key step is substituting the appropriate edge length–radius relationship to eliminate one variable. Each unit cell type yields a characteristic packing efficiency:
| Unit Cell Type | Atoms Per Cell | Edge–Radius Relation | Packing Efficiency |
|---|---|---|---|
| Simple Cubic | 1 | a = 2r | 52% |
| Body-Centered Cubic | 2 | √3 · a = 4r | 68% |
| Face-Centered Cubic | 4 | √2 · a = 4r | 74% |
The Trend to Remember
Packing efficiency increases from SC (52%) to BCC (68%) to FCC (74%). The face-centered cubic structure packs atoms the most efficiently of the three cubic unit cells.
What Is Close Packing of Spheres?
Beyond cubic unit cells, chemists describe crystal structures in terms of how spherical atoms stack in layers. Close packing refers to arrangements that minimize empty space, and there are two principal types: hexagonal close-packed (HCP) and cubic close-packed (CCP). Both close-packed structures achieve a packing efficiency of 74%, the same as the face-centered cubic unit cell.
In any single layer, each atom is surrounded by six nearest neighbors arranged in a hexagonal pattern—this is true for both HCP and CCP. The critical difference lies in how successive layers stack on top of one another.
HCP vs. CCP: 2 Close-Packing Stacking Patterns
When a second layer of atoms is placed on top of the first, the spheres naturally settle into the gaps (or "holes") between first-layer atoms. Not all holes get filled—there are two distinct sets of gaps, and only one set is occupied by the second layer.
The question then becomes: where does the third layer go?
Hexagonal Close-Packed (HCP): ABAB Pattern
In HCP, the third layer sits directly above the first layer, in the same positions. The pattern alternates between just two layer positions: ABABAB...

Cubic Close-Packed (CCP): ABCABC Pattern
In CCP, the third layer occupies the set of holes that neither the first nor the second layer used—a completely new position. The pattern cycles through three distinct positions before repeating: ABCABC... Importantly, the cubic close-packed arrangement is identical to the face-centered cubic (FCC) unit cell.
| Property | Hexagonal Close-Packed (HCP) | Cubic Close-Packed (CCP) |
|---|---|---|
| Stacking Pattern | ABABAB... | ABCABC... |
| Repeating Layers | 2 distinct positions | 3 distinct positions |
| Nearest Neighbors (per atom) | 6 in same layer | 6 in same layer |
| Common In | Pure metals | Pure metals |
Key Insight
Both HCP and CCP achieve excellent packing efficiency and are widely adopted by pure metals. The sole structural difference is whether the third layer repeats the first (HCP) or introduces an entirely new position (CCP).
How Does Bonding Determine Melting Point?
Solids fall into four categories based on the type of bonding or interactions holding their particles together. The general rule is straightforward: the stronger the bonds or interactions, the higher the melting point. Understanding these categories is central to predicting and comparing the physical properties of crystalline and amorphous solids.
| Solid Type | Held Together By | Typical Melting Point |
|---|---|---|
| Molecular | Intermolecular forces | Low |
| Network Covalent | Covalent bonds throughout lattice | Very high |
| Ionic | Ionic bonds | High |
| Metallic | Metallic bonds | Varies widely |
Why Do Molecular Solids Have Low Melting Points?
Molecular solids are held together not by true chemical bonds but by intermolecular forces—interactions that are far weaker than ionic or covalent bonds. Because these forces are relatively easy to overcome, molecular compounds generally melt at low temperatures.
Examples
- Ice (H2O): Melts at 0°C—relatively high for a molecular compound, thanks to hydrogen bonding.
- Propane (C3H8): Melts at −188°C. At room temperature, propane is not just above its melting point but above its boiling point as well, existing as a gas.
Even water's 0°C melting point looks modest when compared to network covalent and ionic solids, which routinely melt above 1000°C.
Why Do Network Covalent Solids Have Extremely High Melting Points?
Network covalent solids consist of an extended lattice of covalent bonds running throughout the entire structure. Melting one of these solids requires breaking those covalent bonds—a process that demands enormous energy.
Notable Examples
- Diamond (C): Among the highest melting points of any substance.
- Graphite (C): Another allotrope of carbon with an extended covalent network.
- Silicon dioxide / Quartz (SiO2): Melts around 1700°C.
Why So Few?
Network covalent solids are rare. Most substances composed entirely of nonmetals form molecular compounds instead. Diamond, graphite, and quartz are essentially the examples you need to know. If a substance is made of only nonmetals and is not one of these, it is almost certainly a molecular solid.
What Determines the Melting Point of Ionic Solids?
Ionic solids form when metals combine with nonmetals (or polyatomic ions), creating a lattice held together by ionic bonds. The strength of those bonds—measured as lattice energy—directly determines the melting point: higher lattice energy means a higher melting point.
Two factors govern lattice energy:
- Ionic charge: Greater charges produce stronger attractions.
- Ion size: Smaller ions form shorter, stronger bonds.
Comparing Ionic Compounds
Consider LiF, CsF, and Li2O:
- Li2O has the highest melting point because the oxide ion (O2−) carries a −2 charge, producing a much larger lattice energy than the −1 fluoride ion.
- LiF > CsF: Both contain +1/−1 ions, but lithium is much smaller than cesium. Smaller cation → shorter bonds → higher lattice energy → higher melting point (LiF: 845°C vs. CsF: 682°C).
What Affects Melting Points of Metallic Solids?
Metallic solids are held together by metallic bonds—a "sea" of delocalized electrons shared across metal cations. The more electrons available for bonding and the smaller the metal atoms, the stronger the metallic bonds and the higher the melting point.
Main Group Metals
For the alkali metals and alkaline earth metals, the trends are clean:
- Going up a group: Smaller atoms form shorter, stronger metallic bonds → higher melting points.
- Going from Group 1 to Group 2: More valence electrons available for bonding → higher melting points.
Example: Alkali Metal Melting Points
Cesium is so soft it would melt in your hand, while sodium melts at 98°C. As you move up the group to smaller atoms, melting points increase significantly.
Transition Metals: A More Complex Picture
Transition metals complicate the trend. You might expect melting points to rise steadily across a period as valence electrons increase, but the reality is more nuanced.
The metals in the middle of each transition series have the most unpaired electrons, and it is these unpaired electrons that contribute most to metallic bonding. As a result, melting points peak near the center of the d-block and decrease toward both ends.
Watch for the Drop-Off
When the d orbitals become fully filled (zinc, cadmium), those electrons are no longer available for bonding. The result is a dramatic decrease in melting point:
- Nickel: >1400°C → Zinc: 420°C
- Palladium: ~1500°C → Cadmium: 321°C
Beyond the transition metals, post-transition metals (to the right of the metalloid staircase) show increasing melting points as additional valence electrons become available for bonding.

Melting Point Rankings for All 4 Solid Types
As a general guide for exam questions, the four solid types rank as follows:
| Rank | Solid Type | Typical Melting Behavior |
|---|---|---|
| 1 (Highest) | Network Covalent | Covalent bonds throughout the lattice → extremely high melting points |
| 2 | Ionic | Strong ionic bonds → high melting points (close second) |
| 3 | Metallic | Broad range—from very high (Fe, Pd ~1500°C) to very low (Cs melts in your hand) |
| 4 (Lowest) | Molecular | Only intermolecular forces → generally the lowest melting points |
Don't Forget the Exceptions
Mercury is a metal that is liquid at room temperature. Ionic liquids exist (though they are extremely rare). Some molecular solids melt at higher temperatures than certain metals. The general ranking holds for most exam scenarios, but be ready for curveball questions about exceptions.
Key Takeaways
- 3 cubic unit cells exist: simple cubic (atoms at corners only), body-centered cubic (corners + body center), and face-centered cubic (corners + face centers).
- Atoms per unit cell: SC = 1, BCC = 2, FCC = 4. Corner atoms contribute 1/8 each; face-center atoms contribute 1/2; body-center atoms contribute 1.
- Coordination numbers increase from SC (6) to BCC (8) to FCC (12), reflecting increasingly tight packing.
- Edge length–radius relationships depend on where atoms touch: SC along the edge (a = 2r), FCC along the face diagonal (√2a = 4r), BCC along the body diagonal (√3a = 4r).
- Packing efficiency increases from SC (52%) to BCC (68%) to FCC (74%).
- Close packing comes in 2 varieties: hexagonal close-packed (ABAB stacking) and cubic close-packed (ABCABC stacking); CCP is equivalent to the FCC unit cell.
- Melting points depend on bonding type: network covalent > ionic > metallic (varies) > molecular, with the strongest bonds or interactions producing the highest melting points.
- For ionic solids, higher charges and smaller ions increase lattice energy and melting point. For metallic solids, smaller atoms and more bonding electrons increase melting point, with transition metals peaking in the middle of the d-block.